CAS: 108-95-2; Phenol
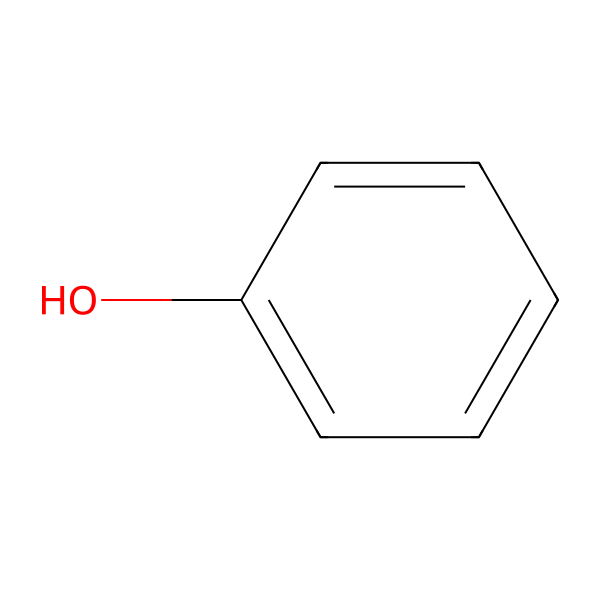
该化合物是一种芳香有机化合物,其特征为氢氧基(-OH)组,与苯环相连,看起来无色至光粉红晶状固体或粘粘液,有明显的甜味,焦油味的味道.Phenol在水,酒精和醚中溶解,并具有酸性和基本特性,使其酸性较弱.其分子配方为C6H6O,与其他许多有机化合物相比,其沸点相对较低.苯醇在塑料,树脂和药物的生产中广泛使用,并且作为各种化学合成物的前体.然而,它也是有毒的,在接触时可引起皮肤刺激和其他健康问题.由于其活性,芬醇可受到电药替代反应,使其在有机合成中成为有价值的化合物.在处理苯酚时,安全防范措施至关重要,因为其腐蚀性及其潜在的环境影响.
- ⚠️危险化学品目录(2015年版)急性毒性-经口,类别3* 《化妆品禁用原料目录》
- 英文名称Phenol
- 中文名称苯酚
- IUPAC名称phenol
- 其它别名Hydroxybenzene
- CAS编号108-95-2
- MFCD编号:MFCD00002143
- EINECS号:203-632-7
- FDA UNII编号:339NCG44TV
- 分子式:C6H6O Lewis structure
- 分子量:94.11
- 产品CID: 125538
- 产品分类原料药 → 人工合成抗感染类药 → 消毒防腐药
- 相似结构搜索
- 产品信息参考
 ;
- InChIKey: ISWSIDIOOBJBQZ-UHFFFAOYSA-N
- InChI=1S/C6H6O/c7-6-4-2-1-3-5-6/h1-5,7H…
- 计算化学: 氢键受体数1.0氢键供体数1.0可旋转键数0.0
物理性质
- 熔点40-42 °C(文献)
- 沸点181.8±0.0 °C at 760 mmHg
- 闪点79.4±0.0 °C
- 密度1.1±0.1 g/cm3
- pKa:9.89(at 20°C)
- PSA:20.23
- LogP:1.48
- 折射率1.553
- 蒸汽压0.6±0.3 mmHg at 25°C
- 溶解性H2O:50 mg/mL at 20 °C,澄清,无色
- 敏感性1.化学性质:苯酚有弱酸性(25°C,Ka=1.3×10-10),与碱作用生成盐。其大多数盐类是水溶性的,能被碳酸(Ka=4.3×10-7)所游离,利用此特性可以区分酚类和羧酸,工业上用来从复杂的煤焦油中分离苯酚。苯酚与氯化铁的水溶液或醇溶液作用。呈现蓝色或紫蓝色。大多数酚类在浓硫酸中加入亚硝酸钾都发生利伯曼(Liebermann)显色反应。苯酚容易发生醚化或酯化反应,可以制得乙酸苯酯(C6H5O°CCH3)、磷酸三苯酯[(C6H5)3PO4]、水杨酸苯酯(C6H5O°C6H5OH)、苯甲醚(C6H5°CH3)、二苯醚(C6H5°C6H5)等。苯酚容易氧化,生成多羟基衍生物、联苯和二苯醚衍生物、醌类、草酸及树脂状物质。苯酚与锌一起蒸馏得到苯,催化加氢生成环己醇。苯酚容易发生亲电取代反应。例如与溴水反应立即生成2,4,6-三溴苯酚,要得到一溴取代物必须用特殊的方法。与稀硝酸在常温即可发生硝化反应,生成邻硝基酚和对硝基酚的混合物。与浓硫酸在常温即可发生磺化反应,主要生成邻羟基苯磺酸,若在100°C下进行磺化,则主要产物为对羟基苯磺酸。苯酚也容易与醇或烯烃发生烷基化反应。与重氮盐发生偶合反应得到偶氮染料。苯酚最主要的反应是与羰基化合物进行缩合,与甲醛水溶液反应生成酚醛树脂,与苯二甲酸酐反应生成酚酞。酚钠与二氧化碳反应生成水杨酸。2.苯酚属高毒类,对皮肤和黏膜有强烈的腐蚀性,又能经皮肤和黏膜吸收而造成中毒,开始出现刺激,局部麻醉,进而变为溃疡。低浓度能使蛋白质变性,高浓度能使蛋白质沉淀,故对各种细胞有直接损害。而且苯酚在体内分离后可造成肾脏损伤,从而引起继发性死亡。误服苯酚时强烈地刺激胃,引起腹部剧痛。与之接触之组织受到明显腐蚀。长期吸入苯酚蒸气时,可患苯酚虚脱症,开始感到头痛、咳嗽、倦怠、虚弱、食欲减退,后期出现不断咳嗽、皮肤痛痒、肾区有压迫感、胸部有沉重感、严重失眠、皮肤苍白、蛋白尿,最后因慢性肾炎而死亡。人口服苯酚的致死量约2~15g,纯苯酚的毒性更大。TJ 36-79规定车间空气中最高容许浓度为5mg/m3。3.稳定性 稳定4.禁配物 强氧化剂、强酸、强碱5.避免接触的条件 光照6.聚合危害 不聚合
- 外观形态透明结晶固体
- 储存条件储存注意事项储存于阴凉、通风的库房。远离火种、热源。避免光照。库温不超过35°C,相对湿度不超过80%。包装密封保存。应与氧化剂、酸类、碱类、食用化学品分开存放,切忌混储。配备相应品种和数量的消防器材。储区应备有合适的材料收容泄漏物。
- 产品应用用作生产酚醛树脂,卡普隆和己二酸的原料,也用于塑料和医药工业.
- 性质描述苯酚为无色针状结晶或白色结晶熔块,可燃,腐蚀力强.有毒.不纯品在光和空气作用下变为淡红或红色,遇碱变色更快.与大约8水混合可液化.可吸收空气中水分并液化.有特殊臭味和燃烧味,极稀的溶液具有甜味.相对密度1.0576,凝固点41°C,熔点43°C,沸点181.7°C(182°C),折射率1.54178,闪点79.44°C(闭杯),85°C(开杯),自燃点715°C,蒸气密度3.24,蒸气压0.13kPa(40.1°C),蒸气与空气混合物燃烧极限1.7-8.6.1g苯酚溶于约15ml水(0.67,25°C加热后可以任何比例溶解),12ml苯.易溶于乙醇,乙醚,氯仿,甘油,二硫化碳,凡士林,挥发油,固定油,强碱水溶液.几乎不溶于石油醚.水溶液pH值约为6.0.
MSDS等安全信息
- GHS象形图


- GHS符号GHS08 & GHS06 & GHS05;
注释: Health hazard & Skull and crossbones & Corrosion - 危险类别致突变性 类别2
急性毒性 类别3(吸入)
急性毒性 类别3(经皮)
急性毒性 类别3(经口)
特定目标器官毒性-重复接触 类别2
皮肤腐蚀 类别1B
水生慢性毒性 类别2
特定目标器官毒性-单次接触 类别1
严重眼损伤 类别1
皮肤腐蚀 类别1
严重眼刺激 类别2
急性毒性 类别1(吸入)
水生急性毒性 类别1
特定目标器官毒性-重复接触 类别1
水生慢性毒性 类别1
生殖毒性 类别1B
Muta. 1B
特定目标器官毒性 - 单次接触 类别3
皮肤腐蚀 类别1A
急性毒性 类别4(吸入)
急性毒性 类别4(经口)
致癌性 类别2
皮肤致敏 类别1
皮肤腐蚀/刺激 类别2
水生慢性毒性 类别3
皮肤腐蚀 类别1C
急性毒性 类别4(经皮)
急性毒性 类别2(吸入) - 警示词Danger(危险)
- 危险描述H341 |怀疑导致遗传缺陷.
H331 |吸入会中毒.
H311 |皮肤接触会中毒.
H301 |吞咽会中毒.
H373 |长期或反复接触可能损害器官.
H314 |造成严重皮肤灼伤和眼损伤.
H411 |对水生生物有毒并具有长期持续影响.
H370 |损害器官.
H318 |造成严重眼损伤.
H319 |造成严重眼刺激.
H330 |吸入致命.
H400 |对水生生物毒性极大.
H372 |长期或反复接触损害器官.
H410 |对水生生物毒性极大并具有长期持续影响.
H360 |可能损害生育能力或胎儿.
H340 |可能导致遗传缺陷.
H335 |可能引起呼吸道刺激.
H332 |吸入有害.
H302 |吞咽有害.
H351 |怀疑致癌.
H317 |可能引起皮肤过敏反应.
H315 |造成皮肤刺激.
H412 |对水生生物有害并具有长期持续影响.
H312 |皮肤接触有害. - 防范说明P203, P260, P261, P262, P264, P270, P271, P280, P301+P316, P301+P330+P331, P302+P352, P302+P361+P354, P304+P340, P305+P354+P338, P316, P318, P319, P321, P330, P361+P364, P363, P403+P233, P405, and P501
- UN编号1671.0
- 危险品标志T:Toxic;C:Corrosive;
- 安全声明S26-S36/37/39-S45-S36/37-S28A-S28-S24/25-S1/2-S36-S16
- 危险类别码R40,R68,R48/20/21/22,R23/24/25,R34
- WGK Germany2
欧盟法规
统一分类与标签压力设备指令-第1组危险流体REACH注册ECHA物质ECHA物质食品接触材料-禁用CMR物质工作场所安全标识要求C&L通报化妆品禁用物质清单CoRAP评估清单REACH预注册废弃物危险特性清单上下游产品
CAS号108-86-1 溴苯 | CAS号106991-88-2 1-[4-[4-[4-(4-a... | CAS号10268-69-6 邻氨基苯甲酸苯酯 | CAS号109392-90-7 苯基吖啶-9-羧酸酯 | CAS号100-67-4 苯酚钾 | CAS号67-56-1 甲醇 | CAS号101-65-5 N,N’-[亚甲基二(4,1-... | CAS号10419-35-9 2-乙氧基苯并二氢吡喃 | CAS号111198-02-8 1-(苯氧基甲基)-1H-苯并三唑 | CAS号108098-38-0 2-[(decylamino)... | CAS号1077-05-0 1,3,2-Dioxaphos...合成工艺路线路线简述
- 合成目标产物 Phenol 主要起始原料 4-Chlorophenol
- (文献来源)合成步骤主要原料 4-Chlorophenol
苯基-Beta-葡萄糖吡喃糖苷置于5 Wt% Ruthenium/carbon,氯化乙酰胆碱,乙二醇体系中,化学反应生成苯酚
参考文献:木质纤维素的氢转移还原催化分馏:单体产率高且选择性可切换
标题:木质纤维素的氢转移还原催化分馏:单体产率高且选择性可切换
摘要:这项工作展示了木质纤维素的氢转移还原催化分馏.通过引入氯化胆碱的氢键受体,可以实现具有可切换选择性的高产率芳香族单体.氯化胆碱极大地改善了木质素的溶解并调整了木质素的氢解作用.所提出的策略促进了有针对性的木质素增值和木质素优先生物精炼厂的发展.
Doi:10.1002/anie.202307116
专利信息
专利号:US-12421534-B2
优先权日:2021-11-10
标 题 :Engineered enzymes and method for the synthesis of diverse tyrosine analogs
发明人:ALMHJELL PATRICK J; ARNOLD FRANCES H
权利人:CALIFORNIA INST OF TECHN
摘要:Provided herein is an engineered tryptophan synthase β-subunit (TrpB) that catalyzes the synthesis of tyrosine, tyrosine analogs, or salts thereof. Also provided herein are methods for preparing tyrosine, tyrosine analogs, or a salt thereof using the engineered TrpB described herein.
专利号:US-2008287649-A1
优先权日:2006-12-29
标 题 :Methods for the synthesis of cyclic peptides
发明人:CHEN LIN; HAN YEUN-KWEI; ROBERTS CHRISTOPHER R
权利人:CHEN LIN; HAN YEUN-KWEI; ROBERTS CHRISTOPHER R
摘要:Methods for the synthesis of cyclic peptides are provided, as well as novel dipeptide compounds. The methods include the solid phase synthesis of a dipeptide, which is the coupled to a second peptide in a solid phase reaction. The peptide is then cyclized following the coupling reaction. The methods and dipeptides are particularly useful for the synthesis of MC-4 receptor agonist peptides.
专利号:US-8138330-B2
优先权日:2006-09-11
标 题 :Process for the synthesis of oligonucleotides
发明人:LEUCK MICHAEL; WOLTER ANDREAS; STUMPE ALFRED
权利人:LEUCK MICHAEL; WOLTER ANDREAS; STUMPE ALFRED; SIGMA ALDRICH CO LLC
摘要:The present invention discloses novel methods for the synthesis of oligonucleotides with nucleoside phosphoramidites on solid supports. The methods comprise the stepwise chain assembly of oligonucleotides on supports with 5′-acyl phosphoramidites. The synthesis cycles consist of a front end deprotection step which is conducted with a solution of a primary amine or a phenolate, a phosphoramidite coupling step with a 5′-acyl nucleoside phosphoramidite in the presence of an activator, a phosphite oxidation step and an optional capping step. The novel methods improve the quality of synthetic oligonucleotides due to the irreversibility of the front end deprotection step, which prevents the formation of deletion sequences, and due to the avoidance of acidic reagents in the synthesis cycles, which prevent the formation of depurination side products. The invention further discloses novel nucleoside phosphoramidite compositions wherein the phosphoramidites carry acyl front end protective groups which are cleavable with primary amines or phenolates. The invention is applicable to the synthesis of oligodeoxyribonucleotides, oligoribonucleotides and oligonucleotides with modifications in their sugar or phosphate groups.
专利号:EP-1634887-A3
优先权日:2004-07-23
标题 :Process for synthesis of complex 2-deoxy-2-iodo pyranosides of high purity, particularly suitable for manufacturing pharmaceutically pure annamycin
发明人:SZEJA WIESLAW; FOKT IZABELA; GRYNKIEWICZ GRZEGORZ
权利人:ZAKLAD BADAWCZO PROD
摘要:The method according to the invention comprises using as glycosyl donors high purity 2-deoxy-2-iodo derivatives of sugars of a defined configuration of C-2 carbon with O-alkyl, S-alkyl, O-aryl, S-aryl, O-heteroaryl, S-heteroaryl, O-acyl, O-silyl, O-N-imido, O-P-(phosphino, phosphono, thiophosphono, selenophosphono) groups at the anomeric position, and subjecting them to a condensation reaction with a compound containing a hydroxyl group or a protected hydroxyl group in the presence of electrophilic reagents, followed by deprotection and isolation of a pharmaceutically pure product. The objective of the present invention is to provide a method for preparing complex glycosides by coupling cyclic multifunctional compounds containing hydroxyl functional groups (glycosyl acceptors, including aglycones of natural origin which are known as constituents of active pharmaceutical ingredients) with diastereoisomerically pure 2-deoxy-2-halopyranosyl derivatives containing anomeric substituent capable of exchange under glycosidating conditions, for example by virtue of activation with electrophilic agents. This method substantially simplifies the synthesis of Annamycin. n The present invention relies on a regio- and stereoselective reaction of cohalogenating 1,2-unsaturated sugars in the presence of hydroxyl group containing compounds, such as: water, alcohols, phenols, carboxylic acids, silanols, phosphinic acids, phosphoric acids, thiophosphoric acids, selenophosphoric acids, followed by separation of the product with proper C-2 carbon configuration required for the synthesis. As the separation of stereoisomers is carried out at the stage of obtaining a relatively inexpensive intermediate in the form of glycosyl donor, the method of the invention substantially reduces the cost of Annamycin manufacture. Other advantages of the synthesis process of the invention consist in the reduction of the number of stages of the antibiotic manufacture process and in yield increase of the final product.
专利号:EP-0671928-B1
优先权日:1992-09-24
标题 :Synthesis of n-substituted oligomers
发明人:ZUCKERMANN RONALD N; KERR JANICE M; KENT STEPHEN BRIAN HENRY; MOOS WALTER H; SIMON REYNA J; GOFF DANE A
权利人:CHIRON CORP
摘要:Poly N-substituted Glycines (poly NSGs), wherein the substituents bear purine or pyrimidine bases (R<9>) every second glycine:
专利号:US-5118853-A
优先权日:1988-10-13
标题:Processes for the synthesis of 3-disubstituted aminoacroleins
发明人:LEE GEORGE T; REPIC OLJAN
权利人:SANDOZ LTD
摘要:Process for the synthesis of compounds of the formula ##STR1## comprising the steps of (i) reacting a compound of the formula ##STR2## with oxalyl chloride or oxalyl bromide to form the corresponding compound of the formula ##STR3## (ii) reacting said compound of the formula ##STR4## with a compound of the formula ##STR5## to form the corresponding compound of the formula ##STR6## (iii) hydrolyzing said compound of the formula ##STR7## to obtain the corresponding compound of the formula ##STR8## the use of the compounds of the formula ##STR9## for the synthesis of the compounds of the formula ##STR10## and the use of the intermediates of Formula VII for the direct synthesis of the compounds of Formula II, n wherein n R 1 is C 1-3 alkyl, phenyl or phenyl substituted by 1 to 3 substituents each of which is independently C 1-3 alkyl, C 1-3 alkoxy, fluoro, chloro, bromo or nitro (maximum of two nitro groups), n R 1b is phenyl or phenyl substituted by 1 to 3 substituents each of which is independently C 1-3 alkyl, C 1-3 alkoxy, fluoro, chloro, bromo or nitro (maximum of two nitro groups), n R 2 is C 1-3 alkyl, n one of R 3 and R 4 is ##STR11## and the other is primary or secondary C 1-6 alkyl not containing an asymmetric carbon atom, C 3-6 cycloalkyl or phenyl-(CH 2 ) m -, n wherein n R 7 is hydrogen, C 1-3 alkyl, n-butyl, i-butyl, t-butyl, C 1-3 alkoxy, n-butoxy, i-butoxy, trifluoromethyl, fluoro, chloro, phenoxy or benzyloxy, n R 8 is hydrogen, C 1-3 alkyl, C 1-3 alkoxy, trifluoromethyl, fluoro, chloro, phenoxy or benzyloxy, n R 9 is hydrogen, C 1-2 alkyl, C 1-2 alkoxy, fluoro or chloro, and n m is 1, 2 or 3, with the provisos that not more than one of R 7 and R 8 is trifluoromethyl, not more than one of R 7 and R 8 is phenoxy, and not more than one of R 7 and R 8 is benzyloxy, n R 5 is hydrogen, C 1-3 alkyl, n-butyl, i-butyl, t-butyl, C 3-6 cycloalkyl, C 1-3 alkoxy, n-butoxy, i-butoxy, trifluoromethyl, fluoro, chloro, phenoxy or benzyloxy, and n R 6 is hydrogen, C 1-3 alkyl, C 1-3 alkoxy, trifluoromethyl, fluoro, chloro, phenoxy, or benzyloxy, with the provisos that not more than one of R 5 and R 6 is trifluoromethyl, not more than one of R 5 and R 6 is phenoxy, and not more than one of R 5 and R 6 is benzyloxy, n R 10 is C 1-6 alkyl, each X is chloro or bromo, and each X.sup.⊖ is chloride or bromide.
网址: http://www.yxzw.com
企业联系电话:0510-87256677;87256688👤
地址:江苏省宜兴市官林镇钮家村
企业联系电话:0510-87256677;87256688👤
📞无锡市展望化工试剂有限公司 ⚠️参考联系方式
联系人:王总
电话:0510-87256677;87256688
手机:18906159966
传真:0510-87256730;87256677
邮箱:sales@yxzw.com
通信地址: 江苏省宜兴市官林镇钮家村
邮编: 214256
电话:0510-87256677;87256688
手机:18906159966
传真:0510-87256730;87256677
邮箱:sales@yxzw.com
通信地址: 江苏省宜兴市官林镇钮家村
邮编: 214256
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:江苏省宜兴市官林镇钮家村
⚠️ 未注册 · 未认证企业
 备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别
✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.sinyear.com
企业联系电话:021-51601195;18930548442👤
地址:上海市闵行区漕宝路1555号商务中心211室
企业联系电话:021-51601195;18930548442👤
📞上海信跃化学品有限公司 ⚠️参考联系方式
联系人:赵巧飞
电话:021-51601195;18930548442
手机:18930548442;18116267768;13524286472
传真:021-51685846
邮箱:645908973@qq.com
通信地址: 上海市闵行区漕宝路1555号商务中心211室
邮编: 200011
电话:021-51601195;18930548442
手机:18930548442;18116267768;13524286472
传真:021-51685846
邮箱:645908973@qq.com
通信地址: 上海市闵行区漕宝路1555号商务中心211室
邮编: 200011
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海市闵行区漕宝路1555号商务中心211室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.qidichem.cn
企业联系电话:18915071511👤
地址:江苏省常州市武进区湖塘镇花园街128-1-16
企业联系电话:18915071511👤
📞常州市启迪化工有限公司 ⚠️参考联系方式
联系人:田经理
电话:18915071511
手机:18915071511
传真:0519-88198608
邮箱:15261185020@139.com
通信地址: 江苏省常州市武进区湖塘镇花园街128-1-16
邮编: 213161
电话:18915071511
手机:18915071511
传真:0519-88198608
邮箱:15261185020@139.com
通信地址: 江苏省常州市武进区湖塘镇花园街128-1-16
邮编: 213161
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:江苏省常州市武进区湖塘镇花园街128-1-16
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.weimingchem.com
企业联系👤
邮箱:sales@weimingchem.com
通信地址: 山东省淄博市齐鲁电商谷
邮编: 255000
地址:山东省淄博市齐鲁电商谷
企业联系👤
📞山东伟明化工有限公司 ⚠️参考联系方式
邮箱:sales@weimingchem.com
通信地址: 山东省淄博市齐鲁电商谷
邮编: 255000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省淄博市齐鲁电商谷
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.jsrchem.cn
企业联系电话:13646408011👤
地址:山东济南市经四路万达广场C座 1515室
企业联系电话:13646408011👤
📞山东金生润化工有限公司 ⚠️参考联系方式
联系人:吕华平
电话:13646408011
邮箱:sales@jsrchem.cn
通信地址: 山东济南市经四路万达广场C座 1515室
邮编: 250001
电话:13646408011
邮箱:sales@jsrchem.cn
通信地址: 山东济南市经四路万达广场C座 1515室
邮编: 250001
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东济南市经四路万达广场C座 1515室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.sdyschem.com
企业联系电话:15069516671👤
地址:山东省济南市天桥区新材料交易中心
企业联系电话:15069516671👤
📞山东宇硕化工有限公司 ⚠️参考联系方式
联系人:李经理
电话:15069516671
邮箱:479104910@qq.com
通信地址: 山东省济南市天桥区新材料交易中心
邮编: 250000
电话:15069516671
邮箱:479104910@qq.com
通信地址: 山东省济南市天桥区新材料交易中心
邮编: 250000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省济南市天桥区新材料交易中心
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.rongyuechem.com
企业联系电话:0533-6288372 13853369850(WhatsApp)👤
地址:山东省淄博市开发区鲁泰大道99号
企业联系电话:0533-6288372 13853369850(WhatsApp)👤
📞山东荣悦化工有限公司 ⚠️参考联系方式
联系人:殷猛
电话:0533-6288372 13853369850(WhatsApp)
手机:13853369850(WhatsApp)
传真:0533-6288182
邮箱:Selina@rongyuechemical.com
通信地址: 山东省淄博市开发区鲁泰大道99号
邮编: 255000
电话:0533-6288372 13853369850(WhatsApp)
手机:13853369850(WhatsApp)
传真:0533-6288182
邮箱:Selina@rongyuechemical.com
通信地址: 山东省淄博市开发区鲁泰大道99号
邮编: 255000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省淄博市开发区鲁泰大道99号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.yidareagent.com
企业联系电话:0579-88766576👤
地址:浙江省兰溪市经济开发区丹溪大道2006号
企业联系电话:0579-88766576👤
📞兰溪市屹达化工试剂有限公司 ⚠️参考联系方式
联系人:朱先生
电话:0579-88766576
手机:13906798056
传真:0579-88766582
邮箱:Sales@yidareagent.com
通信地址: 浙江省兰溪市经济开发区丹溪大道2006号
邮编: 321100
电话:0579-88766576
手机:13906798056
传真:0579-88766582
邮箱:Sales@yidareagent.com
通信地址: 浙江省兰溪市经济开发区丹溪大道2006号
邮编: 321100
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:浙江省兰溪市经济开发区丹溪大道2006号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.zh-chem.com
企业联系电话:0579-88800388👤
地址:浙江省兰溪市凯旋路83号
企业联系电话:0579-88800388👤
📞浙江中星化工试剂有限公司 ⚠️参考联系方式
联系人:程建军
电话:0579-88800388
手机:13967903058
传真:0579-88800388
邮箱:cjj@zh-chem.com
通信地址: 浙江省兰溪市凯旋路83号
邮编: 321100
电话:0579-88800388
手机:13967903058
传真:0579-88800388
邮箱:cjj@zh-chem.com
通信地址: 浙江省兰溪市凯旋路83号
邮编: 321100
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:浙江省兰溪市凯旋路83号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.xingechemical.com
企业联系电话:18669310806👤
地址:山东省临沂市经开区危化品交易市场
企业联系电话:18669310806👤
📞山东辛格化工有限公司 ⚠️参考联系方式
联系人:赵总
电话:18669310806
传真:0539-7127392
邮箱:3216887117@qq.com
通信地址: 山东省临沂市经开区危化品交易市场
邮编: 276000
电话:18669310806
传真:0539-7127392
邮箱:3216887117@qq.com
通信地址: 山东省临沂市经开区危化品交易市场
邮编: 276000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省临沂市经开区危化品交易市场
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.ouyachem.net
企业联系电话:13961201549👤
地址:常州市金坛经济开发区
企业联系电话:13961201549👤
📞常州市金坛欧亚化工有限公司 ⚠️参考联系方式
联系人:欧阳先生
电话:13961201549
手机:13961201549;13814793879
邮箱:sales@ouyachem.net
通信地址: 常州市金坛经济开发区
邮编: 213200
电话:13961201549
手机:13961201549;13814793879
邮箱:sales@ouyachem.net
通信地址: 常州市金坛经济开发区
邮编: 213200
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:常州市金坛经济开发区
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.airui-sh.com
企业联系电话:021-36308843 36308842👤
地址:上海市宝山区富联二路177弄(保集e智谷)4号楼411室
企业联系电话:021-36308843 36308842👤
📞上海艾锐化工有限公司 ⚠️参考联系方式
联系人:张经理
电话:021-36308843 36308842
手机:13611689036
传真:021-36308845
邮箱:121735124@qq.com
通信地址: 上海市宝山区富联二路177弄(保集e智谷)4号楼411室
邮编: 201906
电话:021-36308843 36308842
手机:13611689036
传真:021-36308845
邮箱:121735124@qq.com
通信地址: 上海市宝山区富联二路177弄(保集e智谷)4号楼411室
邮编: 201906
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海市宝山区富联二路177弄(保集e智谷)4号楼411室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.chemdatang.com
企业联系电话:025-85431992👤
地址:南京市玄武区红山路88号常发广场3幢415
企业联系电话:025-85431992👤
📞南京大唐化工有限责任公司 ⚠️参考联系方式
联系人:王梅
电话:025-85431992
手机:13951889902
传真:025-85407165
邮箱:vip@chemdatang.com
通信地址: 南京市玄武区红山路88号常发广场3幢415
邮编: 210000
电话:025-85431992
手机:13951889902
传真:025-85407165
邮箱:vip@chemdatang.com
通信地址: 南京市玄武区红山路88号常发广场3幢415
邮编: 210000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:南京市玄武区红山路88号常发广场3幢415
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.aoxiangchemical.com
企业联系电话:13256141539👤
地址:山东省济南市天桥区济南新材料交易中心
企业联系电话:13256141539👤
📞济南翱翔化工有限公司 ⚠️参考联系方式
联系人:张总
电话:13256141539
邮箱:sales@aoxiangchemical.com
通信地址: 山东省济南市天桥区济南新材料交易中心
邮编: 250000
电话:13256141539
邮箱:sales@aoxiangchemical.com
通信地址: 山东省济南市天桥区济南新材料交易中心
邮编: 250000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省济南市天桥区济南新材料交易中心
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.kuanhao.com
企业联系电话:021-66313386👤
地址:上海静安区江场西路299弄7号1702室
企业联系电话:021-66313386👤
📞上海宽昊工贸有限公司 ⚠️参考联系方式
联系人:吴华
电话:021-66313386
手机:13901628252
传真:021-66313363
邮箱:shkuanhao@163.com
通信地址: 上海静安区江场西路299弄7号1702室
邮编: 200436
电话:021-66313386
手机:13901628252
传真:021-66313363
邮箱:shkuanhao@163.com
通信地址: 上海静安区江场西路299弄7号1702室
邮编: 200436
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海静安区江场西路299弄7号1702室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.tcjxhgc.com
企业联系电话:0536-5717699 5717669👤
地址:山东省寿光市王高工业园
企业联系电话:0536-5717699 5717669👤
📞寿光市天成精细化工厂 ⚠️参考联系方式
联系人:李先生
电话:0536-5717699 5717669
手机:13563608910
传真:0536-5717698
邮箱:lmh@tcjxhgc.com
通信地址: 山东省寿光市王高工业园
邮编: 262709
电话:0536-5717699 5717669
手机:13563608910
传真:0536-5717698
邮箱:lmh@tcjxhgc.com
通信地址: 山东省寿光市王高工业园
邮编: 262709
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省寿光市王高工业园
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
山东昱轩化工产品有限公司免费会员展台
小微企业
信用代码: 91370104MACUTA466P法定代表人:范增建
信用代码: 91370104MACUTA466P法定代表人:范增建
注册资本:300万(元)企业类型:有限责任公司(自然人独资)
成立日期:2023-09-13登记机关:济南市槐荫区市场监督管理局
注册地址: 山东省济南市槐荫区经十路22799号银座中心4号楼,5号楼,裙房5-1408
一般项目:化工产品销售(不含许可类化工产品);石油制品销售(不含危险化学品);气体、液体分离及纯净设备制造;专用化学产品销售(不含危险化学品);金属材料销售;化肥销售;塑料制品销售;国内贸易代理;贸易经纪;货物进出口。(除依法须经批准的项目外,凭营业执照依法自主开展经营活动)
https://www.yxhgcp.com
企业联系电话:18954475867👤
地址: 山东省济南市槐荫区经十路22799号银座中心4号楼,5号楼,裙房5-1408
常规和库存产品*产品总量: 55
企业联系电话:18954475867👤
📞 山东昱轩化工产品有限公司
产品销售经理: 孙经理
邮箱: 776248306@qq.com
联系电话: 18954475867
邮箱: 776248306@qq.com
联系电话: 18954475867
⚠️ 声明: 咨询或交易时请注意风险评估和信息核实,百琢研不参与任何交易。
地址: 山东省济南市槐荫区经十路22799号银座中心4号楼,5号楼,裙房5-1408
常规和库存产品*产品总量: 55
平台资质认证✓营业执照✓
危化品经营许可证
108-95-2
苯酚
苯酚
0.999
包装:200镀锌桶
包装:200镀锌桶
 其他备注: 多年专业化工材料销售服务经验,公司承诺保证产品质量以及供货周期
网址: http://www.hengyuchem.com
企业联系电话:0519-88109093;13915019000👤
地址:常州市钟楼区张家工业园118号
企业联系电话:0519-88109093;13915019000👤
📞常州市恒誉化工有限公司 ⚠️参考联系方式
联系人:黄敏
电话:0519-88109093;13915019000
手机:13915019000
传真:0519-88121023
邮箱:sales@hengyuchem.com
通信地址: 常州市钟楼区张家工业园118号
邮编:
电话:0519-88109093;13915019000
手机:13915019000
传真:0519-88121023
邮箱:sales@hengyuchem.com
通信地址: 常州市钟楼区张家工业园118号
邮编:
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:常州市钟楼区张家工业园118号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.jshyzhg.com
企业联系电话:13952708558👤
地址:江苏省扬州市江都区仙女镇陈甸村
企业联系电话:13952708558👤
📞扬州金石化扬子化工有限公司 ⚠️参考联系方式
联系人:杨益勤
电话:13952708558
手机:13952708558
传真:0514-86836789
邮箱:734171993@qq.com
通信地址: 江苏省扬州市江都区仙女镇陈甸村
邮编: 225200
电话:13952708558
手机:13952708558
传真:0514-86836789
邮箱:734171993@qq.com
通信地址: 江苏省扬州市江都区仙女镇陈甸村
邮编: 225200
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:江苏省扬州市江都区仙女镇陈甸村
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.quantaichem.com
企业联系电话:13605333728 18053390260 18053399982👤
地址:山东省淄博市高新区长征路2甲208号淄博海畅轻工制品有限公司院内综合楼215室
企业联系电话:13605333728 18053390260 18053399982👤
📞山东权泰化工科技有限公司 ⚠️参考联系方式
联系人:耿昌盛
电话:13605333728 18053390260 18053399982
手机:13605333728
邮箱:5419845@qq.com
通信地址: 山东省淄博市高新区长征路2甲208号淄博海畅轻工制品有限公司院内综合楼215室
邮编: 255035
电话:13605333728 18053390260 18053399982
手机:13605333728
邮箱:5419845@qq.com
通信地址: 山东省淄博市高新区长征路2甲208号淄博海畅轻工制品有限公司院内综合楼215室
邮编: 255035
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省淄博市高新区长征路2甲208号淄博海畅轻工制品有限公司院内综合楼215室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.erjiaan.com
企业联系电话:15695486777、0533-5270977👤
地址:山东省淄博市淄川区昆仑镇工业园
企业联系电话:15695486777、0533-5270977👤
📞山东筠晟化工有限公司 ⚠️参考联系方式
联系人:孙经理
电话:15695486777、0533-5270977
手机:15695486777
传真:0533-5270977
邮箱:791770359@qq.com
通信地址: 山东省淄博市淄川区昆仑镇工业园
邮编: 255100
电话:15695486777、0533-5270977
手机:15695486777
传真:0533-5270977
邮箱:791770359@qq.com
通信地址: 山东省淄博市淄川区昆仑镇工业园
邮编: 255100
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省淄博市淄川区昆仑镇工业园
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.sdkaiyue.com
企业联系电话:李经理15265310795👤
电话:李经理15265310795
传真:外贸:徐经理 18253159658
邮箱:sales@sdkaiyue.com
通信地址: 济南市CBD片区华润置地广场6区2号楼6层
邮编: 250100
地址:济南市CBD片区华润置地广场6区2号楼6层
企业联系电话:李经理15265310795👤
📞山东凯越化工股份有限公司 ⚠️参考联系方式
电话:李经理15265310795
传真:外贸:徐经理 18253159658
邮箱:sales@sdkaiyue.com
通信地址: 济南市CBD片区华润置地广场6区2号楼6层
邮编: 250100
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:济南市CBD片区华润置地广场6区2号楼6层
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.jincheng-chem.com
企业联系电话:13906273259;15851296963;15851292163👤
地址:江苏如皋市林梓镇环城北路20号/上海市宝山区城银路318号1509室
企业联系电话:13906273259;15851296963;15851292163👤
📞如皋市天成化工有限公司 ⚠️参考联系方式
联系人:邵俊成
电话:13906273259;15851296963;15851292163
手机:15000741928/13906273259;(邵俊成)15851292163;(邵田明)
传真:0513-81720186 021-66315829/60733796
邮箱:sales@jincheng-chem.com
通信地址: 江苏如皋市林梓镇环城北路20号/上海市宝山区城银路318号1509室
邮编: 226512
电话:13906273259;15851296963;15851292163
手机:15000741928/13906273259;(邵俊成)15851292163;(邵田明)
传真:0513-81720186 021-66315829/60733796
邮箱:sales@jincheng-chem.com
通信地址: 江苏如皋市林梓镇环城北路20号/上海市宝山区城银路318号1509室
邮编: 226512
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:江苏如皋市林梓镇环城北路20号/上海市宝山区城银路318号1509室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.jnhrchem.com
企业联系电话:18560225890👤
地址:山东省济南市天桥区济南新材料新材料产业园区
企业联系电话:18560225890👤
📞济南宏润化工有限公司 ⚠️参考联系方式
联系人:王经理
电话:18560225890
手机:15069195602
传真:0531-68801820
邮箱:sales@jnhrchem.com
通信地址: 山东省济南市天桥区济南新材料新材料产业园区
邮编: 250000
电话:18560225890
手机:15069195602
传真:0531-68801820
邮箱:sales@jnhrchem.com
通信地址: 山东省济南市天桥区济南新材料新材料产业园区
邮编: 250000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省济南市天桥区济南新材料新材料产业园区
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.zizhongchem.com
企业联系电话:13609876053👤
地址:辽宁省沈阳市浑南区全运三路99-1号
企业联系电话:13609876053👤
📞沈阳子仲化工有限公司 ⚠️参考联系方式
联系人:张总
电话:13609876053
手机:13609876053
邮箱:13609876053@139.com
通信地址: 辽宁省沈阳市浑南区全运三路99-1号
邮编:
电话:13609876053
手机:13609876053
邮箱:13609876053@139.com
通信地址: 辽宁省沈阳市浑南区全运三路99-1号
邮编:
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:辽宁省沈阳市浑南区全运三路99-1号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.czhongyu-chem.com
企业联系电话:13813691203👤
地址:江苏省常州市武进区湖塘镇峰云汇花园22幢1422-1423
企业联系电话:13813691203👤
📞常州市弘裕化工有限公司 ⚠️参考联系方式
联系人:袁总
电话:13813691203
手机:13813691203
传真:0519-89996110
邮箱:1031106541@qq.com
通信地址: 江苏省常州市武进区湖塘镇峰云汇花园22幢1422-1423
邮编: 213000
电话:13813691203
手机:13813691203
传真:0519-89996110
邮箱:1031106541@qq.com
通信地址: 江苏省常州市武进区湖塘镇峰云汇花园22幢1422-1423
邮编: 213000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:江苏省常州市武进区湖塘镇峰云汇花园22幢1422-1423
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.shsjchem.com
企业联系电话:15921014380,13167128868👤
地址:上海市宝山区高逸路112-118号
企业联系电话:15921014380,13167128868👤
📞上海珅桔化工有限公司 ⚠️参考联系方式
联系人:蒋经理
电话:15921014380,13167128868
手机:15062602202
传真:13167128868
邮箱:173446357@qq.com
通信地址: 上海市宝山区高逸路112-118号
邮编: 200439
电话:15921014380,13167128868
手机:15062602202
传真:13167128868
邮箱:173446357@qq.com
通信地址: 上海市宝山区高逸路112-118号
邮编: 200439
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海市宝山区高逸路112-118号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.bangchengchem.com
企业联系电话:021-69106990;69106980;13701823733👤
地址:上海市青浦区重固镇赵重公路2278号5号楼3层C57座
企业联系电话:021-69106990;69106980;13701823733👤
📞上海邦成化工有限公司 ⚠️参考联系方式
联系人:卢小姐
电话:021-69106990;69106980;13701823733
手机:13701823733
传真:QQ:89190072
邮箱:13701823733@163.com
通信地址: 上海市青浦区重固镇赵重公路2278号5号楼3层C57座
邮编: 201803
电话:021-69106990;69106980;13701823733
手机:13701823733
传真:QQ:89190072
邮箱:13701823733@163.com
通信地址: 上海市青浦区重固镇赵重公路2278号5号楼3层C57座
邮编: 201803
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海市青浦区重固镇赵重公路2278号5号楼3层C57座
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.njsnchem.com
电话: 025-85525881 13905176698👤
销售电话:025-85525881 13905176698
邮箱:sales@njsnchem.com
电话: 025-85525881 13905176698👤
📞南京三诺化工有限公司 ⚠️参考联系方式
销售电话:025-85525881 13905176698
邮箱:sales@njsnchem.com
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别 ✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。网址: http://www.ksylmy.com
企业联系电话:0512-50366581👤
地址:江苏昆山前进西路1088号虹桥大厦805-807室
企业联系电话:0512-50366581👤
📞昆山市亚龙贸易有限公司 ⚠️参考联系方式
联系人:张经理
电话:0512-50366581
手机:18051886390
传真:0512-57598675
邮箱:ksylmy12@yalongchina.net
通信地址: 江苏昆山前进西路1088号虹桥大厦805-807室
邮编: 215300
电话:0512-50366581
手机:18051886390
传真:0512-57598675
邮箱:ksylmy12@yalongchina.net
通信地址: 江苏昆山前进西路1088号虹桥大厦805-807室
邮编: 215300
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:江苏昆山前进西路1088号虹桥大厦805-807室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.dmreagent.com
企业联系电话:022-24923800;022-24922517👤
地址:天津市东丽区华明镇北于堡村子西
企业联系电话:022-24923800;022-24922517👤
📞天津市大茂化学试剂厂 ⚠️参考联系方式
联系人:李经理
电话:022-24923800;022-24922517
手机:13002254622
传真:022-24921188
邮箱:damao@dmreagent.com
通信地址: 天津市东丽区华明镇北于堡村子西
邮编: 300300
电话:022-24923800;022-24922517
手机:13002254622
传真:022-24921188
邮箱:damao@dmreagent.com
通信地址: 天津市东丽区华明镇北于堡村子西
邮编: 300300
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:天津市东丽区华明镇北于堡村子西
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.sdbmhx.com
企业联系电话:0533-7872088;0533-7872566;郭经理13953343889 韩经理13589577726 徐经理13475339679 张经理15501966313 邢经理13964396037👤
电话:0533-7872088;0533-7872566;郭经理13953343889 韩经理13589577726 徐经理13475339679 张经理15501966313 邢经理13964396037
邮箱:1020308335@qq.com
通信地址: 山东省淄博市高新区中润大道139号
邮编: 255000
地址:山东省淄博市高新区中润大道139号
企业联系电话:0533-7872088;0533-7872566;郭经理13953343889 韩经理13589577726 徐经理13475339679 张经理15501966313 邢经理13964396037👤
📞山东博曼化学有限公司 ⚠️参考联系方式
电话:0533-7872088;0533-7872566;郭经理13953343889 韩经理13589577726 徐经理13475339679 张经理15501966313 邢经理13964396037
邮箱:1020308335@qq.com
通信地址: 山东省淄博市高新区中润大道139号
邮编: 255000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省淄博市高新区中润大道139号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.richjoint.com
企业联系电话:021-22818988👤
地址:上海市普陀区真南路1228号康建商务广场1号楼1502号
企业联系电话:021-22818988👤
📞上海润捷科技发展有限公司 ⚠️参考联系方式
联系人:蔡玮
电话:021-22818988
手机:18017869591
邮箱:caiwei@richjoint.com
通信地址: 上海市普陀区真南路1228号康建商务广场1号楼1502号
邮编: 200331
电话:021-22818988
手机:18017869591
邮箱:caiwei@richjoint.com
通信地址: 上海市普陀区真南路1228号康建商务广场1号楼1502号
邮编: 200331
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海市普陀区真南路1228号康建商务广场1号楼1502号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.lcjinxinda.com
企业联系👤
地址:山东省聊城市高新区
企业联系👤
📞聊城金信达新材料有限公司 ⚠️参考联系方式
联系人:于霞
通信地址: 山东省聊城市高新区
邮编: 252000
通信地址: 山东省聊城市高新区
邮编: 252000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省聊城市高新区
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.shbrchem.com
企业联系电话:13262886496;17625729319👤
地址:上海市宝山区城银路525号1903室
企业联系电话:13262886496;17625729319👤
📞上海宝瑞化工有限公司 ⚠️参考联系方式
联系人:于先生;郑女士
电话:13262886496;17625729319
手机:13262886496;17625729319
传真:021-62148466
邮箱:2425283046@qq.com
通信地址: 上海市宝山区城银路525号1903室
邮编: 200332
电话:13262886496;17625729319
手机:13262886496;17625729319
传真:021-62148466
邮箱:2425283046@qq.com
通信地址: 上海市宝山区城银路525号1903室
邮编: 200332
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海市宝山区城银路525号1903室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.syshiji.net
企业联系电话:024-87375999,87228199👤
地址:辽宁康平经济开发区朝阳工业园
企业联系电话:024-87375999,87228199👤
📞沈阳化学试剂厂 ⚠️参考联系方式
联系人:夏晓光
电话:024-87375999,87228199
手机:13591410607
传真:024-27345668
邮箱:sales@syshiji.net
通信地址: 辽宁康平经济开发区朝阳工业园
邮编: 110032
电话:024-87375999,87228199
手机:13591410607
传真:024-27345668
邮箱:sales@syshiji.net
通信地址: 辽宁康平经济开发区朝阳工业园
邮编: 110032
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:辽宁康平经济开发区朝阳工业园
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.jiubangchem.com.cn
企业联系电话:021-36031288,31169627,31169935👤
地址:上海闸北区江场西路299弄6号楼1003室
企业联系电话:021-36031288,31169627,31169935👤
📞上海九邦化工有限公司 ⚠️参考联系方式
联系人:丁海清
电话:021-36031288,31169627,31169935
手机:13818932992
传真:021-66315995
邮箱:jiubang_168@163.com
通信地址: 上海闸北区江场西路299弄6号楼1003室
邮编: 200436
电话:021-36031288,31169627,31169935
手机:13818932992
传真:021-66315995
邮箱:jiubang_168@163.com
通信地址: 上海闸北区江场西路299弄6号楼1003室
邮编: 200436
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海闸北区江场西路299弄6号楼1003室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://zghysz.webh.testwebsite.cn
企业联系电话:13065055261👤
地址:济南市天桥区华山西路8号
企业联系电话:13065055261👤
📞山东满堂红新材料有限公司 ⚠️参考联系方式
联系人:王琳
电话:13065055261
手机:13065055261
邮箱:1643589804@qq.com
通信地址: 济南市天桥区华山西路8号
邮编: 250000
电话:13065055261
手机:13065055261
邮箱:1643589804@qq.com
通信地址: 济南市天桥区华山西路8号
邮编: 250000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:济南市天桥区华山西路8号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.beierchem.cn
企业联系电话:0570-5663486👤
地址:浙江省衢州市常山县天马街道白龙路10幢西单元602室
企业联系电话:0570-5663486👤
📞常山贝尔化学有限公司 ⚠️参考联系方式
联系人:朱经理
电话:0570-5663486
手机:13706703335
传真:0570-5663486
邮箱:sales@beierchem.cn
通信地址: 浙江省衢州市常山县天马街道白龙路10幢西单元602室
邮编:
电话:0570-5663486
手机:13706703335
传真:0570-5663486
邮箱:sales@beierchem.cn
通信地址: 浙江省衢州市常山县天马街道白龙路10幢西单元602室
邮编:
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:浙江省衢州市常山县天马街道白龙路10幢西单元602室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.shtychem.com
企业联系电话:021-69186366;69182866;13701855675👤
地址:上海市青浦区重固镇赵重公路2278号5号楼3层E100座
企业联系电话:021-69186366;69182866;13701855675👤
📞上海彤源化工有限公司 ⚠️参考联系方式
联系人:单小姐
电话:021-69186366;69182866;13701855675
手机:13701855675
传真:QQ:89093626
邮箱:tongyuanchem@126.com
通信地址: 上海市青浦区重固镇赵重公路2278号5号楼3层E100座
邮编: 201803
电话:021-69186366;69182866;13701855675
手机:13701855675
传真:QQ:89093626
邮箱:tongyuanchem@126.com
通信地址: 上海市青浦区重固镇赵重公路2278号5号楼3层E100座
邮编: 201803
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海市青浦区重固镇赵重公路2278号5号楼3层E100座
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.jych.com
企业联系电话:0519-86666676👤
地址:常州市钟楼区西林街道张家村118号1号楼303室
企业联系电话:0519-86666676👤
📞常州市佳业化工有限公司 ⚠️参考联系方式
联系人:巢依桐
电话:0519-86666676
手机:18661144123
传真:0519-86699676
邮箱:1661209034@qq.com
通信地址: 常州市钟楼区西林街道张家村118号1号楼303室
邮编: 213042
电话:0519-86666676
手机:18661144123
传真:0519-86699676
邮箱:1661209034@qq.com
通信地址: 常州市钟楼区西林街道张家村118号1号楼303室
邮编: 213042
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:常州市钟楼区西林街道张家村118号1号楼303室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.jjlchem.com
企业联系电话:021-52915085;021-52910829;15021318527👤
地址:上海中山北路2052号13楼
企业联系电话:021-52915085;021-52910829;15021318527👤
📞上海金锦乐实业有限公司 ⚠️参考联系方式
联系人:黄小姐
电话:021-52915085;021-52910829;15021318527
手机:15021318527
传真:021-52917765
邮箱:jjlchem@jjlchem.com
通信地址: 上海中山北路2052号13楼
邮编: 200063
电话:021-52915085;021-52910829;15021318527
手机:15021318527
传真:021-52917765
邮箱:jjlchem@jjlchem.com
通信地址: 上海中山北路2052号13楼
邮编: 200063
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海中山北路2052号13楼
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.longxingchem.cn
企业联系电话:0546-6893053/6896098👤
地址:山东省东营市广饶县大王经济开发区
企业联系电话:0546-6893053/6896098👤
📞东营市隆兴化工有限责任公司 ⚠️参考联系方式
联系人:田秀华
电话:0546-6893053/6896098
手机:13561012177
传真:0546-6896098
邮箱:longxinghg@126.com
通信地址: 山东省东营市广饶县大王经济开发区
邮编: 257335
电话:0546-6893053/6896098
手机:13561012177
传真:0546-6896098
邮箱:longxinghg@126.com
通信地址: 山东省东营市广饶县大王经济开发区
邮编: 257335
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省东营市广饶县大王经济开发区
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.yishengdachem.com
企业联系电话:0943-8224312👤
地址:甘肃省白银市兰州路16号
企业联系电话:0943-8224312👤
📞白银易盛达化工有限公司 ⚠️参考联系方式
联系人:魏总
电话:0943-8224312
手机:13893019895
邮箱:sales@jingyouchem.com
通信地址: 甘肃省白银市兰州路16号
邮编: 730900
电话:0943-8224312
手机:13893019895
邮箱:sales@jingyouchem.com
通信地址: 甘肃省白银市兰州路16号
邮编: 730900
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:甘肃省白银市兰州路16号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.denuochem.com
企业联系电话:021-62142081👤
地址:上海中山北路2000号
企业联系电话:021-62142081👤
📞上海德诺化工有限公司 ⚠️参考联系方式
联系人:胡寅卿
电话:021-62142081
手机:13816783342
传真:021-62142081
邮箱:denuochem@163.com
通信地址: 上海中山北路2000号
邮编: 200061
电话:021-62142081
手机:13816783342
传真:021-62142081
邮箱:denuochem@163.com
通信地址: 上海中山北路2000号
邮编: 200061
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海中山北路2000号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.xiechen.com
企业联系电话:021-52908009👤
地址:上海市中山北路2000号中期大厦1659室
企业联系电话:021-52908009👤
📞上海协辰进出口有限公司 ⚠️参考联系方式
联系人:陈经理
电话:021-52908009
手机:13918039333
传真:021-52902009
邮箱:2691179155@qq.com
通信地址: 上海市中山北路2000号中期大厦1659室
邮编: 200072
电话:021-52908009
手机:13918039333
传真:021-52902009
邮箱:2691179155@qq.com
通信地址: 上海市中山北路2000号中期大厦1659室
邮编: 200072
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海市中山北路2000号中期大厦1659室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.youpengchem.com
企业联系电话:13701776567;021-69002118;69006118👤
地址:上海市嘉定区鹤旋路26弄22号512室
企业联系电话:13701776567;021-69002118;69006118👤
📞上海有朋化工有限公司 ⚠️参考联系方式
联系人:傅小姐
电话:13701776567;021-69002118;69006118
传真:QQ:2880675224
邮箱:youpengchem@163.com
通信地址: 上海市嘉定区鹤旋路26弄22号512室
邮编: 201803
电话:13701776567;021-69002118;69006118
传真:QQ:2880675224
邮箱:youpengchem@163.com
通信地址: 上海市嘉定区鹤旋路26弄22号512室
邮编: 201803
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海市嘉定区鹤旋路26弄22号512室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.luohengchem.com
企业联系电话:18953927278(微信同号);16510354444👤
地址:山东聊城东昌府区光岳北路与财干路交叉口东南角别墅区51栋
企业联系电话:18953927278(微信同号);16510354444👤
📞山东洛恒化工产品有限公司 ⚠️参考联系方式
联系人:成经理
电话:18953927278(微信同号);16510354444
手机:18953927278;16510354444
传真:QQ:412139239
邮箱:412139239@qq.com
通信地址: 山东聊城东昌府区光岳北路与财干路交叉口东南角别墅区51栋
邮编:
电话:18953927278(微信同号);16510354444
手机:18953927278;16510354444
传真:QQ:412139239
邮箱:412139239@qq.com
通信地址: 山东聊城东昌府区光岳北路与财干路交叉口东南角别墅区51栋
邮编:
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东聊城东昌府区光岳北路与财干路交叉口东南角别墅区51栋
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.heyichem.com.cn
企业联系电话:021-66310288👤
地址:上海闸北区灵石路709号55号楼C2101室
企业联系电话:021-66310288👤
📞上海和逸化工有限公司 ⚠️参考联系方式
联系人:杨建
电话:021-66310288
手机:13611862688
传真:021-66313635
邮箱:shheyihuagong@163.com
通信地址: 上海闸北区灵石路709号55号楼C2101室
邮编: 200072
电话:021-66310288
手机:13611862688
传真:021-66313635
邮箱:shheyihuagong@163.com
通信地址: 上海闸北区灵石路709号55号楼C2101室
邮编: 200072
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海闸北区灵石路709号55号楼C2101室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.njwchem.com
企业联系电话:021-39127857;39127857;39125966👤
地址:上海市嘉定区南翔镇科福路378号
企业联系电话:021-39127857;39127857;39125966👤
📞上海南佶威实业有限公司 ⚠️参考联系方式
联系人:何经理
电话:021-39127857;39127857;39125966
手机:13003264939
传真:021-69927276
邮箱:sales@njwchem.com
通信地址: 上海市嘉定区南翔镇科福路378号
邮编: 201802
电话:021-39127857;39127857;39125966
手机:13003264939
传真:021-69927276
邮箱:sales@njwchem.com
通信地址: 上海市嘉定区南翔镇科福路378号
邮编: 201802
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:上海市嘉定区南翔镇科福路378号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.qzhychem.com
企业联系电话:13305701112👤
地址:浙江省衢州市柯城区花园明盛小区9-18号
企业联系电话:13305701112👤
📞衢州市华亚化工有限公司 ⚠️参考联系方式
联系人:程先生
电话:13305701112
手机:13305701112
传真:0570-3091487
邮箱:sales@qzhychem.com
通信地址: 浙江省衢州市柯城区花园明盛小区9-18号
邮编: 324004
电话:13305701112
手机:13305701112
传真:0570-3091487
邮箱:sales@qzhychem.com
通信地址: 浙江省衢州市柯城区花园明盛小区9-18号
邮编: 324004
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:浙江省衢州市柯城区花园明盛小区9-18号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.nanlinchem.cn
企业联系电话:0533-3177201 13505332357👤
地址:山东省淄博市高新区汇银国际1104室
企业联系电话:0533-3177201 13505332357👤
📞南林(山东)新材料科技有限公司 ⚠️参考联系方式
联系人:荆先生
电话:0533-3177201 13505332357
手机:13505332357
传真:0533-3585201
邮箱:yetti@163.com
通信地址: 山东省淄博市高新区汇银国际1104室
邮编: 255000
电话:0533-3177201 13505332357
手机:13505332357
传真:0533-3585201
邮箱:yetti@163.com
通信地址: 山东省淄博市高新区汇银国际1104室
邮编: 255000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:山东省淄博市高新区汇银国际1104室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.czyubochem.com
企业联系电话:丁经理15295086982;张经理13961462977👤
地址:江苏省常州市钟楼区万泰国际广场21楼2110室
企业联系电话:丁经理15295086982;张经理13961462977👤
📞常州宇博化工科技有限公司 ⚠️参考联系方式
联系人:丁益平;张经理
电话:丁经理15295086982;张经理13961462977
手机:15295086982;13961462977
传真:0519-88368866
邮箱:2360549695@qq.com
通信地址: 江苏省常州市钟楼区万泰国际广场21楼2110室
邮编:
电话:丁经理15295086982;张经理13961462977
手机:15295086982;13961462977
传真:0519-88368866
邮箱:2360549695@qq.com
通信地址: 江苏省常州市钟楼区万泰国际广场21楼2110室
邮编:
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:江苏省常州市钟楼区万泰国际广场21楼2110室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.ristchem.com
企业联系电话:0712-3185660👤
地址:湖北省孝感市应城市东马坊东城工业园2号
企业联系电话:0712-3185660👤
📞湖北瑞斯特化学试剂有限公司 ⚠️参考联系方式
联系人:林先生
电话:0712-3185660
手机:13476869296
传真:-
邮箱:sales@ristchem.com
通信地址: 湖北省孝感市应城市东马坊东城工业园2号
邮编:
电话:0712-3185660
手机:13476869296
传真:-
邮箱:sales@ristchem.com
通信地址: 湖北省孝感市应城市东马坊东城工业园2号
邮编:
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:湖北省孝感市应城市东马坊东城工业园2号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.liaida.com.cn
企业联系电话:0510-86817326;86802659;86815605👤
地址:江苏省江阴市澄江中路5-1号东都国际大厦603室
企业联系电话:0510-86817326;86802659;86815605👤
📞江阴市利爱达贸易有限公司 ⚠️参考联系方式
联系人:徐顾军;徐金林
电话:0510-86817326;86802659;86815605
手机:13961625969;13906162756
传真:0510-86817326
邮箱:jyliaida@163.com
通信地址: 江苏省江阴市澄江中路5-1号东都国际大厦603室
邮编: 214431
电话:0510-86817326;86802659;86815605
手机:13961625969;13906162756
传真:0510-86817326
邮箱:jyliaida@163.com
通信地址: 江苏省江阴市澄江中路5-1号东都国际大厦603室
邮编: 214431
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:江苏省江阴市澄江中路5-1号东都国际大厦603室
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.puhuachem.com
企业联系电话:0519-83519199;83519308;83519358👤
地址:常州市新北区罗溪镇塑化路25-8号105
企业联系电话:0519-83519199;83519308;83519358👤
📞常州市普华化工科技有限公司 ⚠️参考联系方式
联系人:周玉正
电话:0519-83519199;83519308;83519358
手机:13906122795
传真:0519-88861308
邮箱:sales@puhuachem.com
通信地址: 常州市新北区罗溪镇塑化路25-8号105
邮编:
电话:0519-83519199;83519308;83519358
手机:13906122795
传真:0519-88861308
邮箱:sales@puhuachem.com
通信地址: 常州市新北区罗溪镇塑化路25-8号105
邮编:
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:常州市新北区罗溪镇塑化路25-8号105
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.ziboyixinchem.com
企业联系电话:0533-2833223 13506432153👤
地址:淄博张店共青团西路95号钻石商务大厦18层O户
企业联系电话:0533-2833223 13506432153👤
📞淄博益新化工有限公司 ⚠️参考联系方式
联系人:魏馥荣
电话:0533-2833223 13506432153
手机:13506432153
传真:0533-2869262
邮箱:sales@ziboyixinchem.com
通信地址: 淄博张店共青团西路95号钻石商务大厦18层O户
邮编: 255000
电话:0533-2833223 13506432153
手机:13506432153
传真:0533-2869262
邮箱:sales@ziboyixinchem.com
通信地址: 淄博张店共青团西路95号钻石商务大厦18层O户
邮编: 255000
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:淄博张店共青团西路95号钻石商务大厦18层O户
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
网址: http://www.shsfhx.com
企业联系电话:021-31261285/13391235085👤
地址:宝山区市台路515弄51号
企业联系电话:021-31261285/13391235085👤
📞上海森斐化学品有限公司 ⚠️参考联系方式
联系人:郭冰
电话:021-31261285/13391235085
手机:13391235085
传真:021-31263390
邮箱:16967351@qq.com
通信地址: 宝山区市台路515弄51号
邮编: 200444
电话:021-31261285/13391235085
手机:13391235085
传真:021-31263390
邮箱:16967351@qq.com
通信地址: 宝山区市台路515弄51号
邮编: 200444
🆔 联系时候可告知是从"百琢研"平台获取的信息.
⚠️ 声明: 该企业未认证、未认领,请自行辨别信息的真实性和可靠性。咨询或交易时请注意风险评估与信息核实,百琢研不参与任何交易。企业认领注册入口→
地址:宝山区市台路515弄51号
⚠️ 未注册 · 未认证企业
备注: 📌 数据来源说明:本展台内容基于各搜索引擎等公开数据整理,仅作展示用途。请用户自行辨别✉️ 若企业需抹除展台内容或有异议, 请通过页面底部联系方式告知我们,我们会尽快处理。
第 1 / 1 页
供应商参考报价(招募中)
品牌试剂参考报价(招募中)
📌 第三方产品分析报告
主要参考文献
参考标题:Alkyl 1-Chloroalkyl Carbonates: Reagents For The Synthesis Of Carbamates And Protection Of Amino Groups作者:Gérard Barcelo,Jean-Pierre Senet,Gérard Sennyey,Jean Bensoam,Albert Loffet
摘要:The Synthesis Of 1-Chloroalkyl Carbonates And Their Reaction With Various Type Of Amines Are Described. This Reaction Is Useful For The Synthesis Of Carbamate Pesticides And For The Protection Of Various Amino Groups, Including Amino Acids.
合成参考文献
摘要:HANSCH,C ET AL. (1995)
参考文献:10.1007/bf00174117
摘要:Achterrath-Tuckermann U, Saano V, Minker E, Stroman F, Arny I, Joki S, Nuutinen J, Szelenyi I. Influence of azelastine and some selected drugs on mucociliary clearance. Lung. 1992;170(4):201–9. doi: 10.1007/bf00174117.
参考文献:10.1046/j.1523-1747.1999.00756.x
摘要:Yang F, Brown TL, Cornelius J, Babcock GF, Das PK, Boissy RE, Le Poole IC. Altered Gene Expression in Melanocytes Exposed to 4-Tertiary Butyl Phenol (4-TBP): Upregulation of the A2b Adenosine Receptor1. Journal of Investigative Dermatology. 1999 Nov;113(5):725–31. doi: 10.1046/j.1523-1747.1999.00756.x.
📝 需求与反馈
尽可能描述清楚需求与问题信息